CCS evidence has to stay current.Not just ready for inspection.

INSUS consolidates monitoring, qualification and batch record evidence against EU GMP Annex 1 requirements, and flags approaching deadlines and gaps for the quality team to close. Quality teams keep every determination and sign-off.

  • 01

    It tracks, checks and flags. It never self-certifies.

  • 02

    No monitoring or testing is performed. Only records are read.

  • 03

    Quality signs off before anything enters the system of record.

The problem

A numerically specific regulation,evidenced across scattered systems.

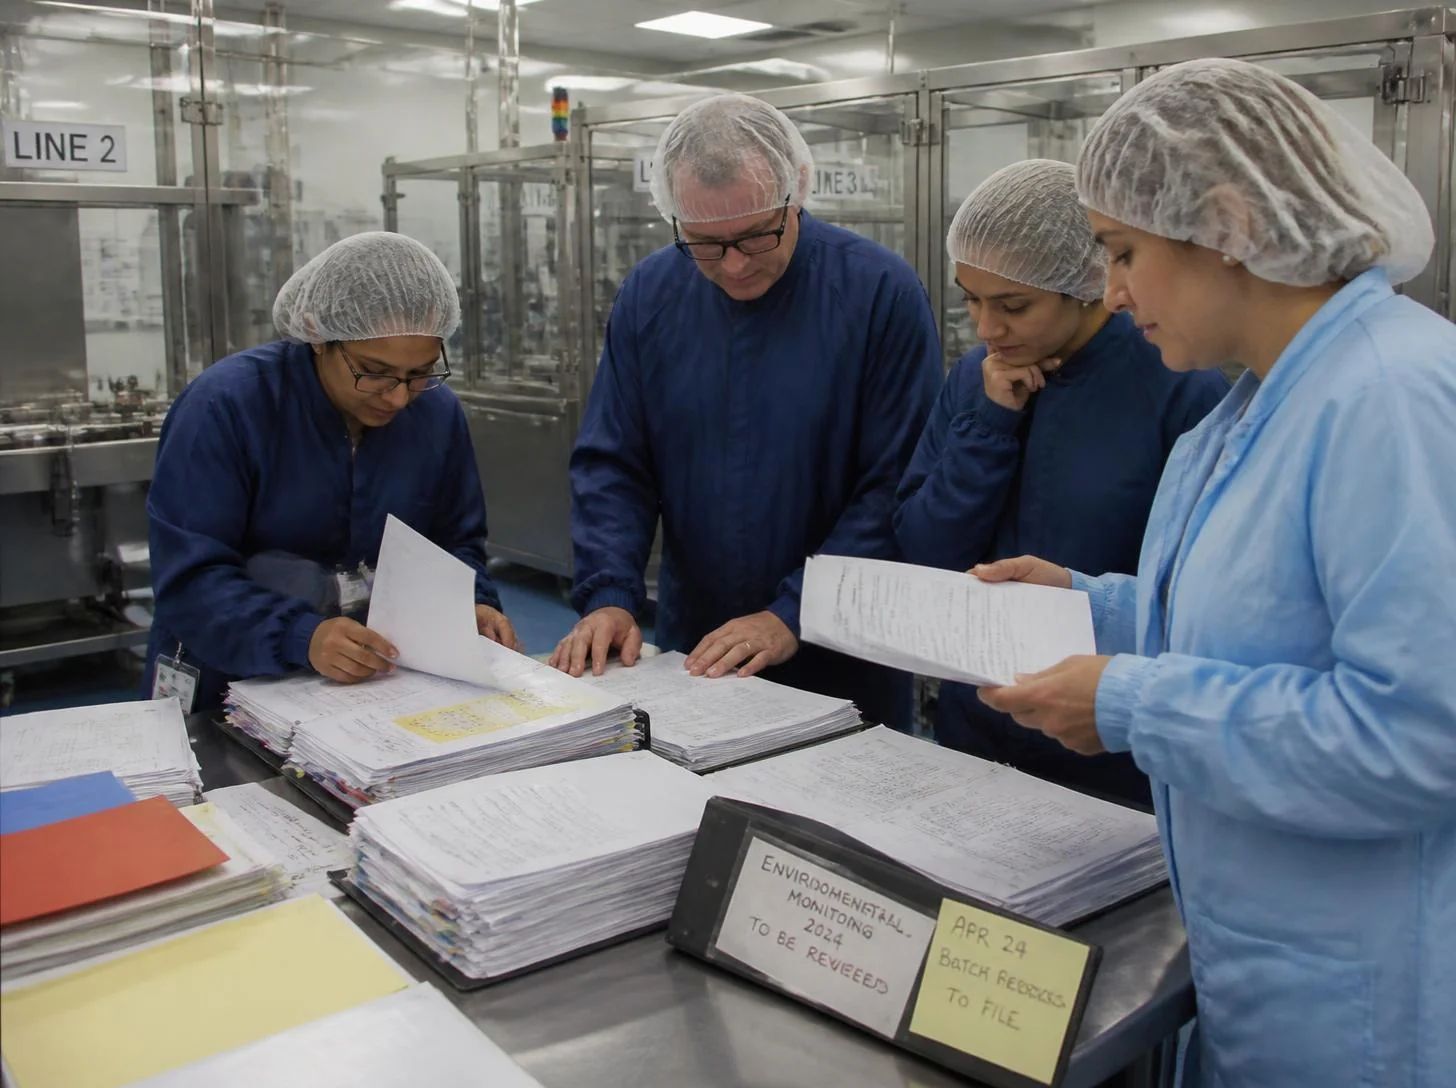
Annex 1 sets grade-dependent particle and microbial limits, guidance values such as differential pressure between rooms, and named frequencies for cleanroom requalification, sterilisation revalidation and gowning reassessment. It also expects interventions, sterilisation cycles and hold times to be documented in the batch record. A Contamination Control Strategy must show ongoing conformance, yet the evidence often sits in separate systems. Approaching deadlines and documentation gaps are frequently found close to a review or inspection.

Sterile manufacturing cleanroom where contamination control evidence originates
01
02
03
  • 01Monitoring by room and grade
  • 02Result trending toward limit
  • 03Requalification approaching

The shift

From evidence assembled for the inspectionto evidence that stays current.

Quality judgement and sign-off stay where they are. What changes is whether evidence is checked continuously against named requirements instead of rebuilt reactively.

Today

Evidence compiled when someone asks for it

  • Monitoring results, requalification dates and revalidation records sit in separate systems.
  • Approaching deadlines are often found close to, or after, the date.
  • Batch record gaps may only appear during batch record review.
With INSUS

Evidence checked against named requirements

  • Monitoring and qualification records are checked against grade limits and named frequencies.
  • Approaching deadlines can be flagged with lead time to act.
  • Documentation gaps can be surfaced before they become review findings.

How it works

From monitoring and qualification recordsto a tracked evidence base.

Delivered through ExpertFlow AI as document intelligence over contamination control records. It organizes and checks evidence. It does not perform environmental monitoring, qualification testing or certify compliance.

  1. Connect

    Bring the evidence into one view

    Connect existing environmental monitoring data, requalification and revalidation records, QMS and MES or batch records including intervention logs and sterilisation cycle identifiers, hold-time records and the current Annex 1 text.

  2. Understand

    Map records to grade-specific requirements

    Map monitoring and qualification records against the grade-dependent limits and named frequencies that apply to each room, grade and piece of equipment.

  3. Identify

    Flag deadlines, trends and documentation gaps

    Surface approaching requalification and revalidation deadlines, monitoring results trending toward a limit, and batch record gaps such as a missing intervention log or incomplete hold-time record.

  4. Advise

    Rank actions for the quality team

    Present a prioritized list of documentation and scheduling actions ranked by deadline proximity and regulatory significance, with evidence mapped to each named control element for review.

  5. Verify

    Quality review before anything enters the record

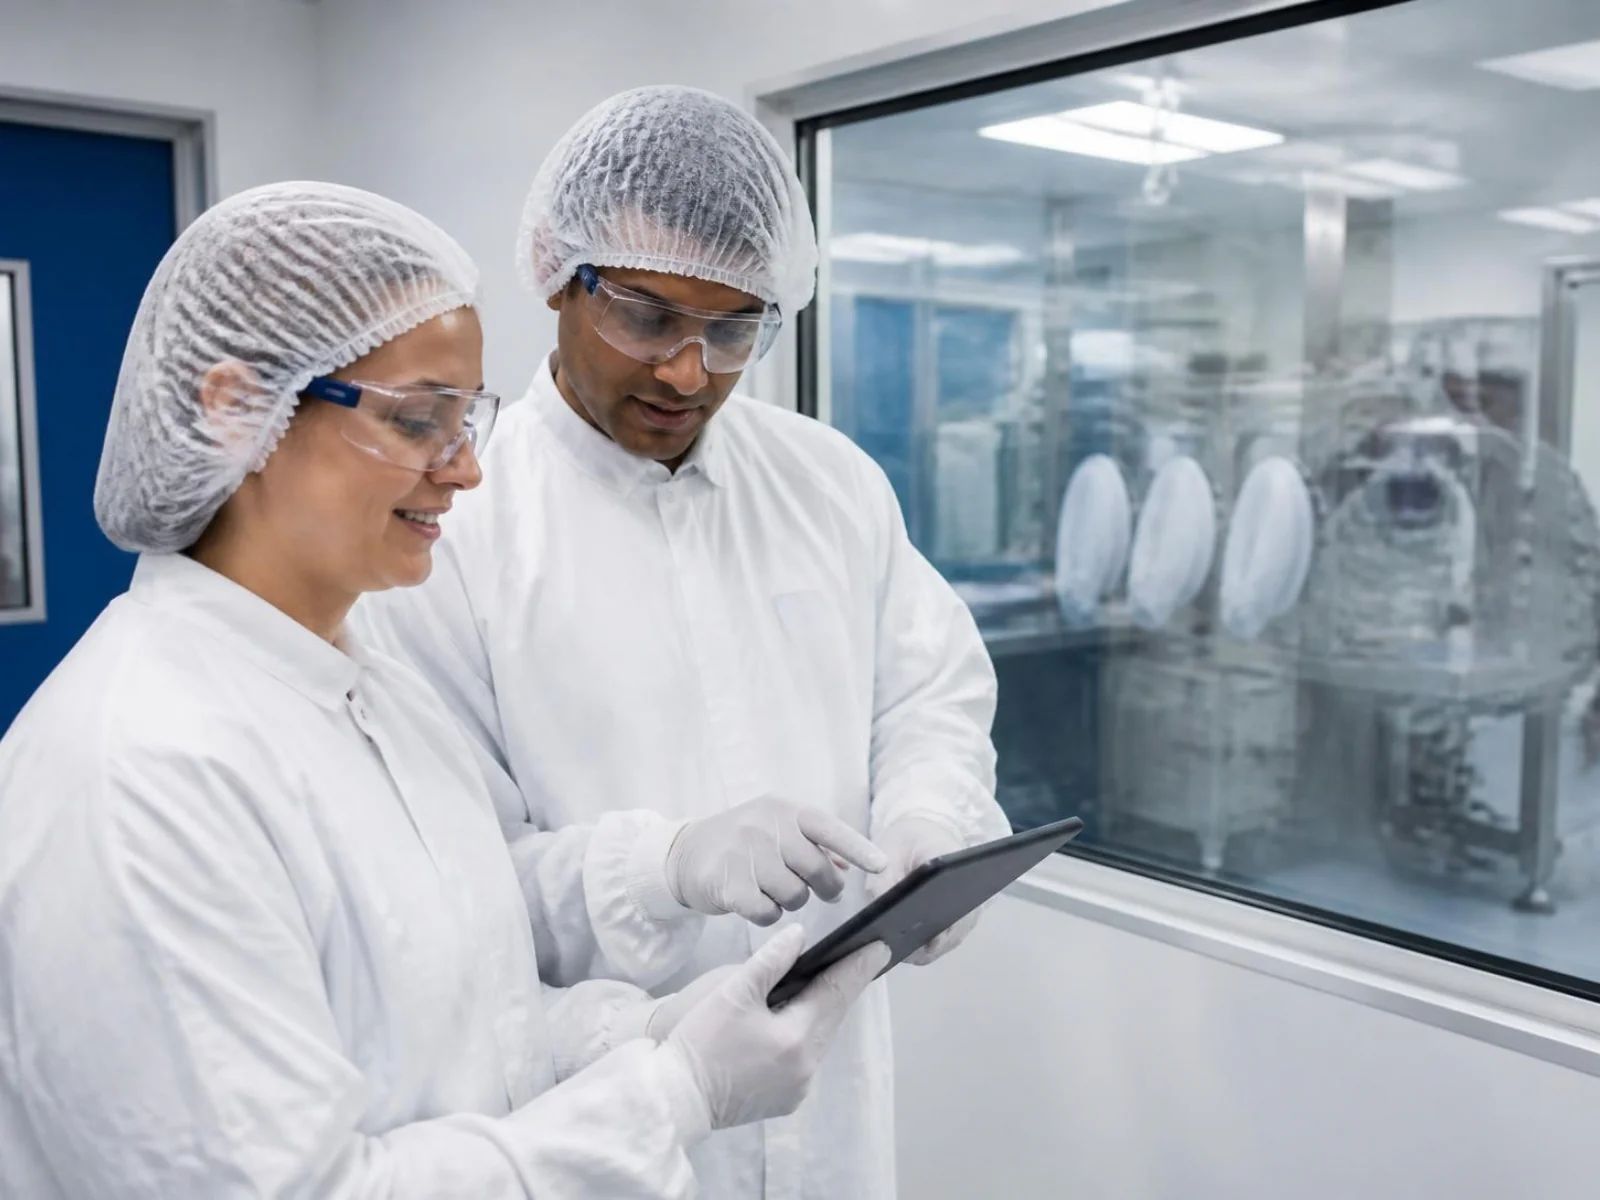
    Quality teams review and sign off every assembled documentation update and flagged trend before it becomes part of the system of record.

Quality team reviewing contamination control evidence before sign-off

Getting started

Start with one sterile suiteand the records you already keep.

For sterile biologics and cell and gene therapy quality teams. Delivered through ExpertFlow AI as document intelligence and deadline tracking. Expert Services can help with discovery and validation. Advisory only. No new sensors. Does not touch plant control systems.

  • What we need. Existing environmental monitoring results, requalification and revalidation records and schedules, hold-time records and batch record documentation for the rooms and equipment in scope. No new hardware.
  • What you see first. One sterile manufacturing suite: evidence checked against named requirements, plus a prioritized list of approaching deadlines and documentation gaps for quality-team review.
  • Where the limits are. It can only check records that exist and that it can reach. It performs no monitoring or testing and does not certify compliance, release a batch or schedule qualification activity on its own.
Explore ExpertFlow AI

Questions quality teams ask us

Annex 1 requires continuous contamination control evidence across monitoring, requalification and batch records. This use case consolidates that evidence, checks it against named requirements and flags gaps for quality-team review.

Existing monitoring results, requalification and revalidation records, hold-time records and batch documentation for rooms and equipment in scope. No new sensors. It organizes records from testing you already perform.

No. It does not predict physical outcomes. It retrieves and links records, tracks deadlines, analyzes monitoring trends in documentation terms and flags gaps for human review.

No. It has no authority to certify conformance, release a batch or independently schedule qualification or validation activity.

No. The quality team reviews and signs off every update and owns every compliance and scheduling decision. Integration is with quality, batch record and qualification systems, not plant control systems.

Through ExpertFlow AI as document intelligence over contamination control records. Expert Services can support discovery, integration and validation with your quality team.

Usefulness depends on accessible underlying records. The system flags gaps for human review; it does not make regulatory determinations or replace GMP procedures.

Biotech · EU GMP Annex 1

Stop rebuilding the evidence before every inspection.

Start with one sterile manufacturing suite. The useful first answer is which deadlines and documentation gaps are already close, while there is still lead time to act on them.